


Abstract
The thus-far unknown ureic derivative dimethyl 2,2′-[carbonylbis(azanediyl)](2S,2′S)-bis[3-(4-hydroxyphenyl)propanoate] has been efficiently synthesized by enantiospecific oxidative carbonylation of readily available l-tyrosine methyl ester, using a very simple catalytic system (PdI2 in conjunction with KI) under relatively mild conditions (100 °C for 5 h in DME as the solvent and under 20 atm of a 4:1 mixture CO-air).
1. Introduction
Ureas are very important carbonyl compounds. Many ureic derivatives have shown interesting biological activities, including anticancer activity [1,2]. Moreover, they find different practical applications, including their use as gelators or hydrogen-bond donors [3,4].
A very attractive method for the preparation of ureas is based on direct carbonylation of amines under oxidative conditions [5,6]. In this field, we have previously reported that a very simple catalytic system, consisting of PdI2 in conjunction with KI, is able to promote the oxidative carbonylation of primary amines to symmetrically 1,2-disubstituted ureas as well as of primary and secondary amines to trisubstituted ureas with excellent selectivities (up to 99%) and very high turnover numbers (up to 43,500 mmol of urea per mmol of palladium) [7,8,9].
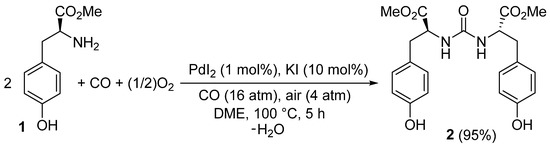
In this Short Note, we report the application of our method to the enantiospecific synthesis of the previously unknown 2′-[carbonylbis(azanediyl)](2S,2′S)-bis[3-(4-hydroxyphenyl)propanoate] 2 starting from commercially available l-tyrosine methyl ester, according to Scheme 1.
Scheme 1.
Enantiospecific synthesis of 2′-[carbonylbis(azanediyl)](2S,2′S)-bis[3-(4-hydroxyphenyl)propanoate] 2 by PdI2/KI-catalyzed oxidative carbonylation of commercially available l-tyrosine methyl ester 1.
2. Results
2′-[Carbonylbis(azanediyl)](2S,2'S)-bis[3-(4-hydroxyphenyl)propanoate] 2 was prepared by enantiospecific palladium-catalyzed oxidative carbonylation of commercially available l-tyrosine methyl ester 1, under the following reaction conditions (Scheme 1): PdI2, 1 mol %; KI, 10 mol %; solvent, 1,2-dimethoxyethane (DME; substrate concentration: 1 mmol per mL of DME); T = 100 °C; P = 20 atm (at 25 °C) of a 4:1 mixture of CO-air. The desired product was obtained in 95% isolated yield after 5 h reaction time and purification of the crude mixture by column chromatography on silica gel (eluent: hexane-AcOEt from 1:1 to 4:6), and was fully characterized by LC-MS spectrometry, IR, 1H-NMR and 13C spectroscopies, elemental analysis, and specific optical rotation.
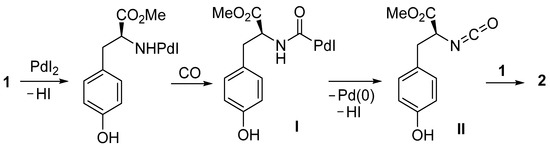
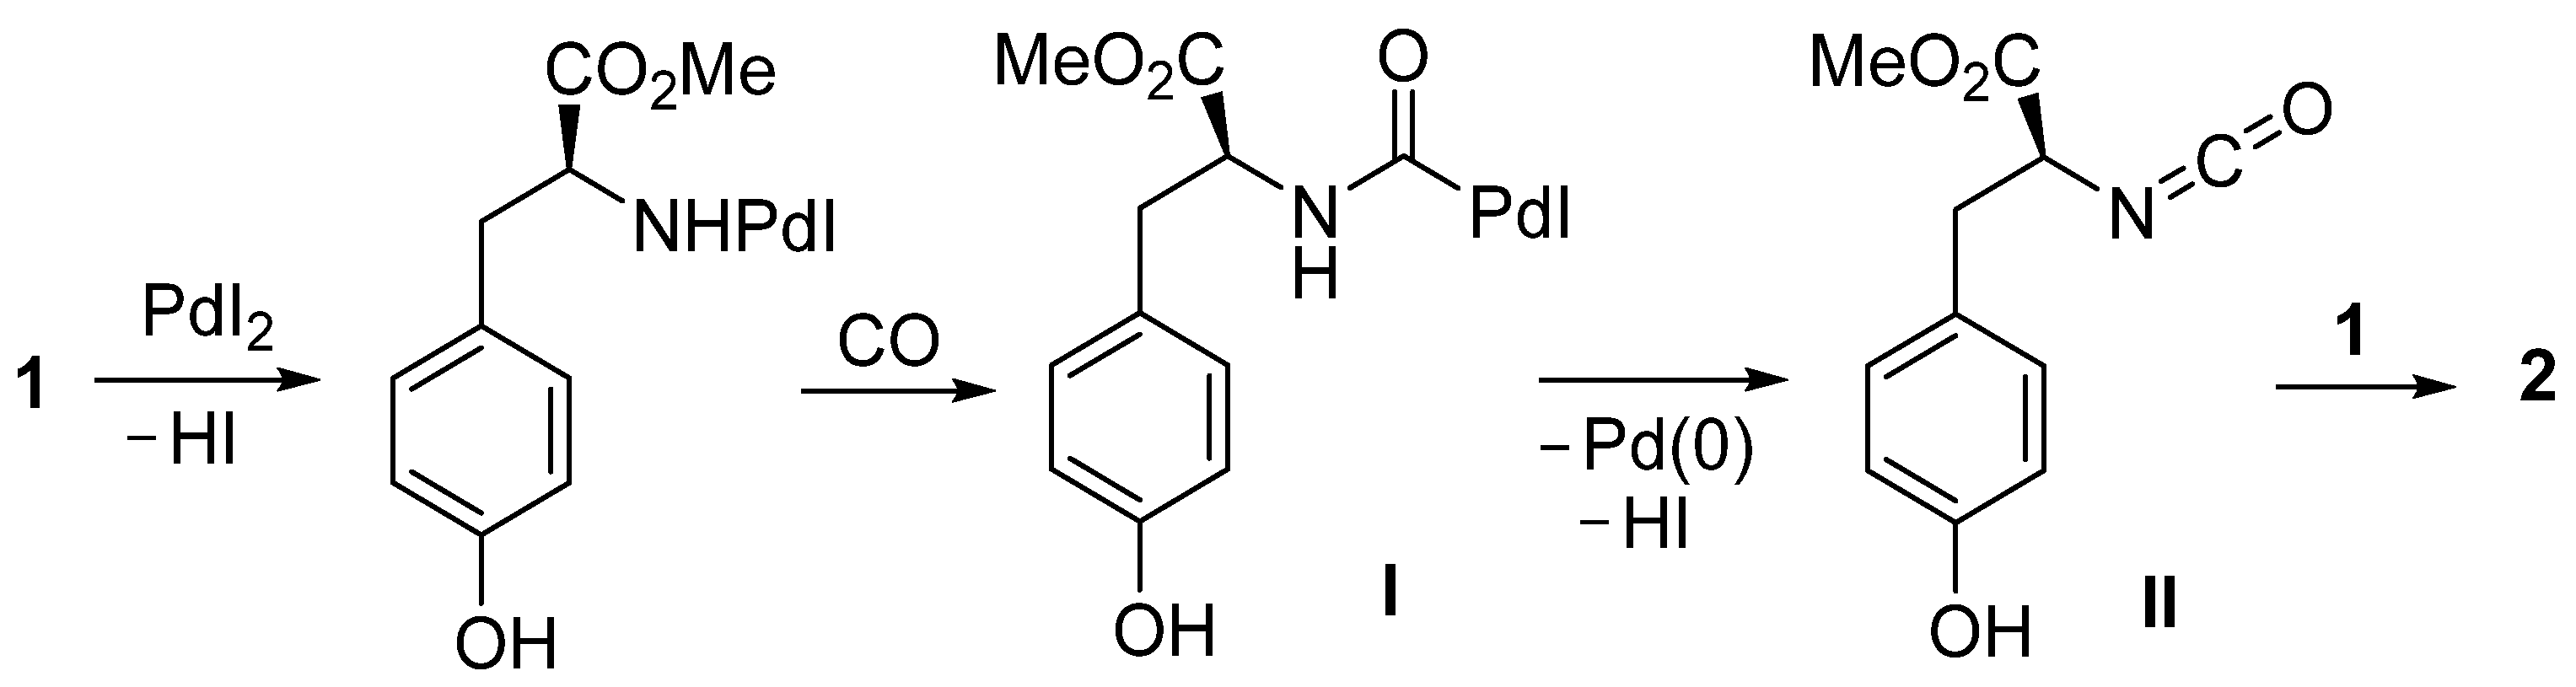
According to our previous findings, the formation of urea 2 can be interpreted as occurring as shown in Scheme 2, involving the formation of a carbamoylpalladium iodide intermediate I (by the reaction between the amino group of the substrate, CO, and PdI2) followed by β-H elimination from the Pd-(CO)-NH moiety to give isocyanate II, and nucleophilic addition to the latter by a second molecule of substrate.
Scheme 2.
Proposed mechanism for the formation of urea 2 from l-tyrosine methyl ester 1.
3. Materials and Methods
Solvents and chemicals were reagent grade and used without further purification. Reactions were analyzed by thin layer chromatography (TLC) on silica gel 60 F254 (Merck s.p.a., Vimodrone, Milano, Italy). Starting material l-tyrosine methyl ester was commercially available (Sigma-Aldrich Italia s.r.l., Milano, Italy). Column chromatography was performed on silica gel 60 (Merck s.p.a., Vimodrone, Milano, Italy, 70−230 mesh). Evaporation refers to the removal of solvent under reduced pressure. Melting point is uncorrected. 1H-NMR and 13C-NMR spectra were recorded at 25 °C on a 300 MHz spectrometer (Bruker DPX Avance 300, Bruker Italia s.r.l., Milano, Italy) in DMSO-d6 solutions with Me4Si as the internal standard. Chemical shifts (δ) and coupling constants (J) are given in ppm and Hz, respectively. IR spectrum was taken with a JASCO FTIR 4200 spectrometer. Mass spectrum was obtained using a HPLC/ESI/Q-TOF HRMS apparatus. HPLC conditions were as follows: water, acetonitrile, and formic acid were of HPLC/MS grade; the HPLC system was an Agilent 1260 Infinity; a reversed-phase C18 column (ZORBAX Extended-C18 2.1 × 50 mm, 1.8 μm) with a Phenomenex C18 security guard column (4 mm × 3 mm) were used; the flow-rate was 0.4 mL/min and the column temperature was set to 30 °C; the eluents were formic acid–water (0.1:99.9, v/v) (phase A) and formic acid–acetonitrile (0.1:99.9, v/v) (phase B); the following gradient was employed: 0–10 min, linear gradient from 5% to 95% B; 10–15 min, washing and reconditioning of the column to 5% B; injection volume was 10 μL; the eluate was monitored through MS TIC. The mass spectra was recorded using an Agilent 6540 UHD accurate-mass Q-TOF spectrometer equipped with a Dual AJS ESI source working in negative mode, under the following conditions: N2 was employed as desolvation gas at 300 °C and a flow rate of 9 L/min; the nebulizer was set to 45 psig; the sheath gas temperature was set at 400 °C and a flow of 12 L/min; a potential of 2.7 kV was used on the capillary for positive ion mode; the fragmentor was set to 175 V; the MS spectrum was recorded in the 150–1000 m/z range. Microanalysis was carried out in our analytical laboratory (Thermo–Fischer Elemental Analyzer Flash 2000, Rodano, Milano, Italy).
Synthetic procedure for the preparation of 2′-[carbonylbis(azanediyl)](2S,2′S)-bis[3-(4-hydroxyphenyl)propanoate] 2. A 250 mL stainless steel autoclave was charged in the presence of air with PdI2 (8.2mg, 0.023 mmol), KI (36.5 mg, 0.22 mmol), and a solution of l-tyrosine methyl ester 1 (433.6 mg, 2.22 mmol) in DME (2.2 mL). While stirring, the autoclave was pressurized with CO (16 atm) and air (up to 20 atm) and then heated at 100 °C for 5 h. After cooling, the autoclave was degassed and solvent evaporated. Some amount of crude product was already present in suspension in the reaction mixture. Crude urea 2 was further purified by column chromatography on silica gel using hexane/ethyl acetate from 1/1 to 4/6 as eluent. Yield: 439.3 mg, 95% based on starting 1. Pure 2′-[carbonylbis(azanediyl)](2S,2′S)-bis[3-(4-hydroxyphenyl)propanoate] 2 was a pale yellow solid, m.p. = 89–91 °C. (acetone, c = 6.51 × 10⁻³ g·mL−1) = +215.1°; IR (KBr): ν = 3389 (m, br), 1728 (s), 1651 (w), 1557 (m), 1514 (m), 1443 (m), 1368 (w), 1219 (s), 1115 (w), 841 (w), 802 (m) cm–1; 1H-NMR (300 MHz, DMSO-d6): δ = 9.29 (s, 2H, 2 OH), 6.97–6.90 (m, 4H, aromatic), 6.71–6.63 (m, 4H, aromatic), 6.44 (d, J = 7.9, 2H, 2 NH), 4.34–4.23 (m, 2H, 2 CHCH2), 3.58 (s, 6H, 2 CO2Me), 2.89–2.70 (m, 4H, 2 CHCH2); 13C-NMR (75 MHz, DMSO-d6): δ = 172.8, 156.6, 156.0, 130.0, 126.7, 115.0, 54.2, 51.6, 36.6; LC-MS: m/z = 415 [(M − H)ˉ]; anal. calcd for C21H24N2O7 (416.43): C, 60.88; H, 6.10; N, 6.87; found: C, 60.86; H, 6.13; N, 6.85. Copies of the 1H-NMR and 13C-NMR spectra are given in the Supplementary Materials.
Supplementary Materials
1H-NMR and 13C-NMR spectra for product 2 are available online at http://www.mdpi.com/1422-8599/2018/1/M983/s1.
Acknowledgments
The work was supported by the University of Calabria.
Author Contributions
B.G., R.M. and N.D.C. conceived and designed the experiments; K.F.G. and R.M. performed the experiments; C.C. analyzed and confirmed the data analysis; A.P.P. contributed analysis tools and performed the HPLC-MS experiments; B.G. wrote the paper.
Conflicts of Interest
The authors declare no conflict of interest.
References
- Li, H.-Q.; Lv, P.-C.; Yan, T.; Zhu, H.-L. Urea derivatives as anticancer agents. Anti-Cancer Agents Med. Chem. 2009, 9, 471–480. [Google Scholar] [CrossRef] [Scilit]
- Jagtap, A.D.; Kondekar, N.B.; Sadani, A.A.; Chern, J.-W. Ureas: applications in drug design. Curr. Med. Chem. 2017, 24, 622–651. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yamanaka, M. Urea derivatives as low-molecular-weight gelators. J. Incl. Phenom. Macrocycl. Chem. 2013, 77, 33–48. [Google Scholar] [CrossRef] [Scilit]
- Volz, N.; Clayden, J. The urea renaissance. Angew. Chem. Int. Ed. 2011, 50, 12148–12155. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Beller, M. (Ed.) Catalytic Carbonylation Reactions; Springer: Berlin, Germany, 2006; ISBN 978-3-540-33003-5. [Google Scholar]
- Diaz, D.J.; Darko, A.K.; McElwee-White, L. Transition metal-catalyzed oxidative carbonylation of amines to ureas. Eur. J. Org. Chem. 2007, 2007, 4453–4465. [Google Scholar] [CrossRef] [Scilit]
- Gabriele, B.; Mancuso, R.; Salerno, G.; Costa, M. A novel and efficient method for the Pd-catalysed oxidative carbonylation of amines to symmetrically and unsymmetrically substituted ureas. Chem. Commun. 2003, 2003, 467–487. [Google Scholar] [CrossRef] [Scilit]
- Gabriele, B.; Salerno, G.; Mancuso, R.; Costa, M. Efficient synthesis of ureas by direct palladium-catalyzed oxidative carbonylation of amines. J. Org. Chem. 2004, 69, 4741–4750. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Della Ca’, N.; Bottarelli, P.; Dibenedetto, A.; Aresta, M.; Gabriele, B.; Salerno, G.; Costa, M. Palladium-catalyzed synthesis of symmetrical urea derivatives by oxidative carbonylation of primary amines in carbon dioxide medium. J. Catal. 2011, 282, 120–127. [Google Scholar] [CrossRef] [Scilit]
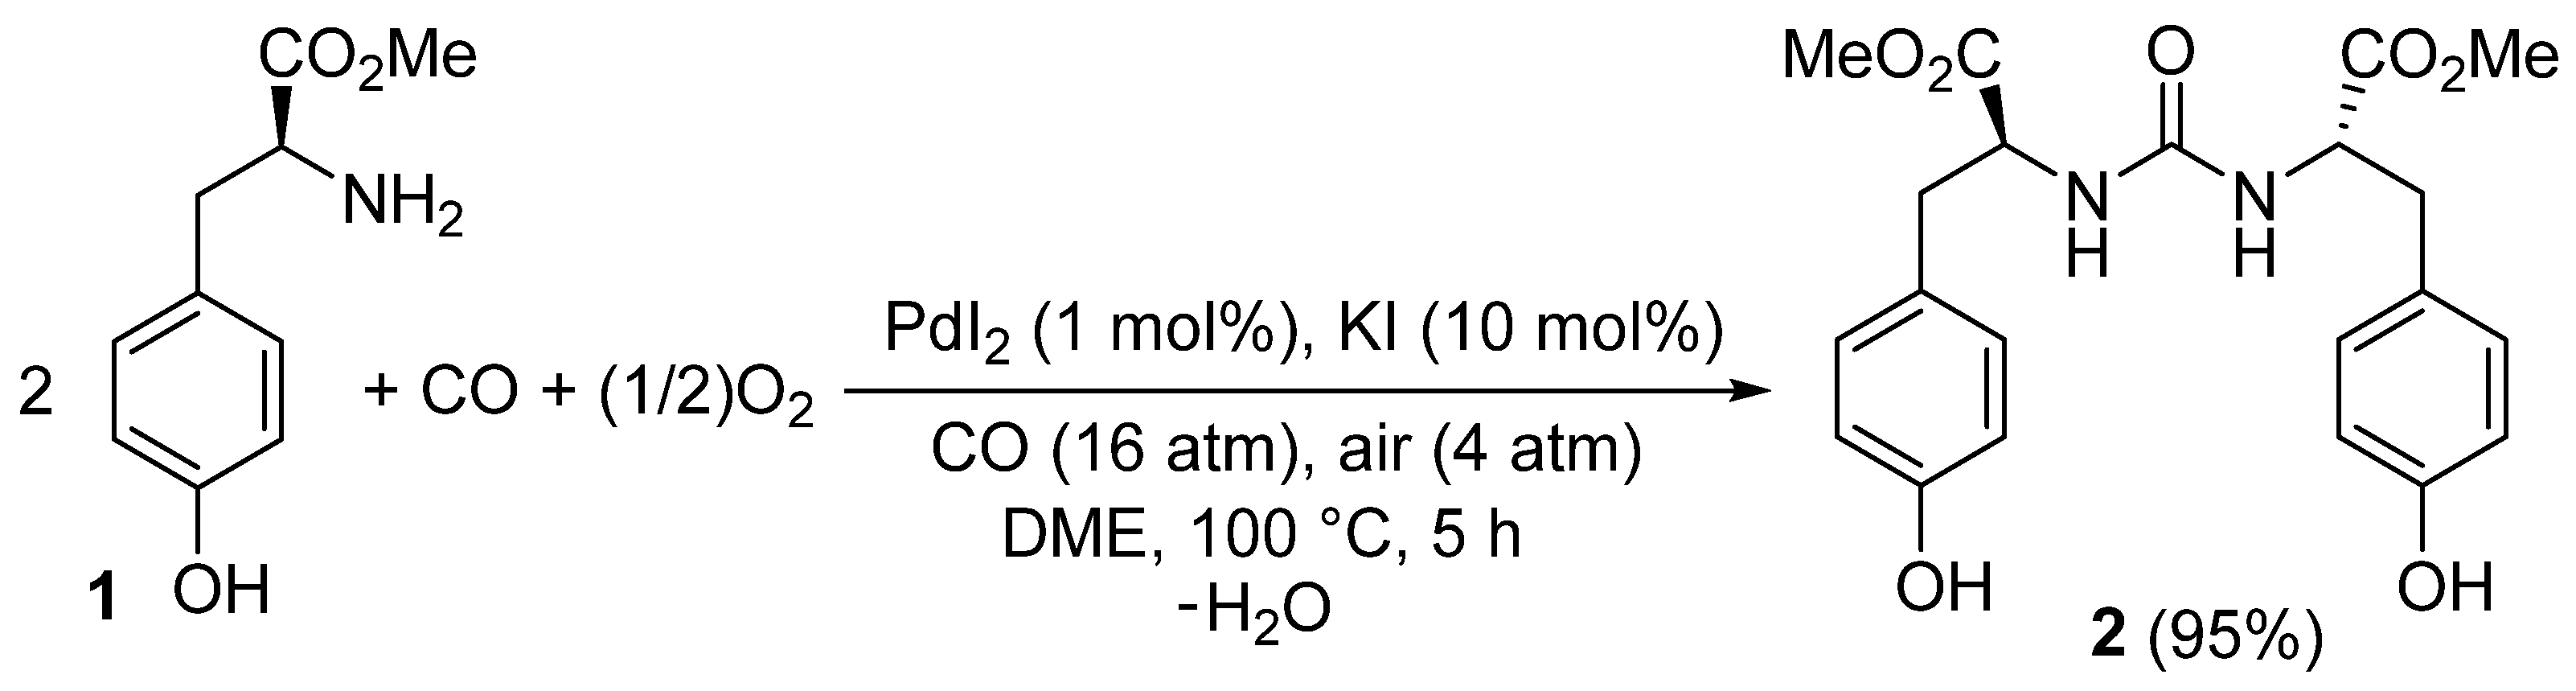
© 2018 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (http://creativecommons.org/licenses/by/4.0/).